The architecture, current capabilities, and recommended use of MACS3 is described, providing an updated reference for applying the MACS framework in contemporary bulk and single-cell regulatory genomics workflows.
Abstract
Since the original publication of Model-based Analysis for ChIP-Seq (MACS), the software has been widely used to identify enriched genomic regions in ChIP-seq, ATAC-seq, CUT&RUN, DNase-seq, and related regulatory genomics assays. Over the years, MACS has evolved substantially, with MACS version 3 (MACS3) now serving as the actively maintained implementation. MACS3 preserves the core MACS framework for fragment pileup, dynamic local background noise, statistical enrichment testing, and peak refinement, while adding functionality needed for contemporary bulk and single-cell workflows. It supports conventional bulk peak calling, paired-end and fragment-based file formats, modular signal processing, direct analysis of single-cell ATAC-seq fragment files, barcode-restricted pseudobulk and cluster-level peak calling, specialized ATAC-seq and variant-calling modules, as well as command-line and programmatic interfaces. MACS3 is distributed through standard software channels and supported by continuous testing across operating systems, Python versions, and CPU architectures. Here we describe the architecture, current capabilities, and recommended use of MACS3, providing an updated reference for applying the MACS framework in contemporary bulk and single-cell regulatory genomics workflows. MACS3 is open-source software available at https://github.com/macs3-project/MACS.
Bulk RNA-seq and single-cell RNA-seq (scRNA-seq) are widely used to investigate gene-expression changes, but downstream analysis often requires multiple statistical, visualization, and reporting tools, creating fragmented workflows that are difficult to configure and reproduce. We developed CoTRA (Comprehensive Toolbox...
Background/Objectives: ATAC-seq footprinting can infer transcription-factor (TF) occupancy across the genome at near-base-pair resolution. However, its broad adoption is limited by high computational demands, complex command-line workflows, and fragmented support for bulk data with biological replicates and single-cell...
The results indicate that GmGM provides a unified, reproducible framework for joint cell clustering and gene-network inference, capable of revealing cellular structure beyond that captured by conventional pipelines.
O. Lanzetta, L. Cutillo, Bailey Andrew et al.· 0 citations
FetchPA is an intuitive pipeline which allows users with virtually no scripting and version control experience to install and manage all software for end-to-end analyses of ATAC-Seq data and guides the user through the identification of differentially accessible regions.
Dustin R. Fetch, Alexey A. Soshnev· bioRxiv· 0 citations
Malva is presented, a computational platform that enables ultrafast, species-agnostic and reference-free interrogation of the raw sequence space, enabling searching for any sequence, mutation, splice junction or pathogen, or spatial location of arbitrary transcripts.
D. León-Periñán, Nikos Karaiskos, N. Rajewsky· Nature· 1 citation
EISCA and EISTA, two standardized, end-to-end pipelines for single-cell RNA-seq and imaging-based spatial transcriptomics analysis, built on the Nextflow nf-core framework provide efficient, flexible, and scalable solutions for comprehensive single-cell and spatial transcriptomics analyses.
Hui-Hai Wu, Ashleigh Lister, Iain C. Macaulay et al.· bioRxiv· 0 citations
Related blog posts
MIT News · Artificial Intelligence· news.mit.eduOct 6, 2026
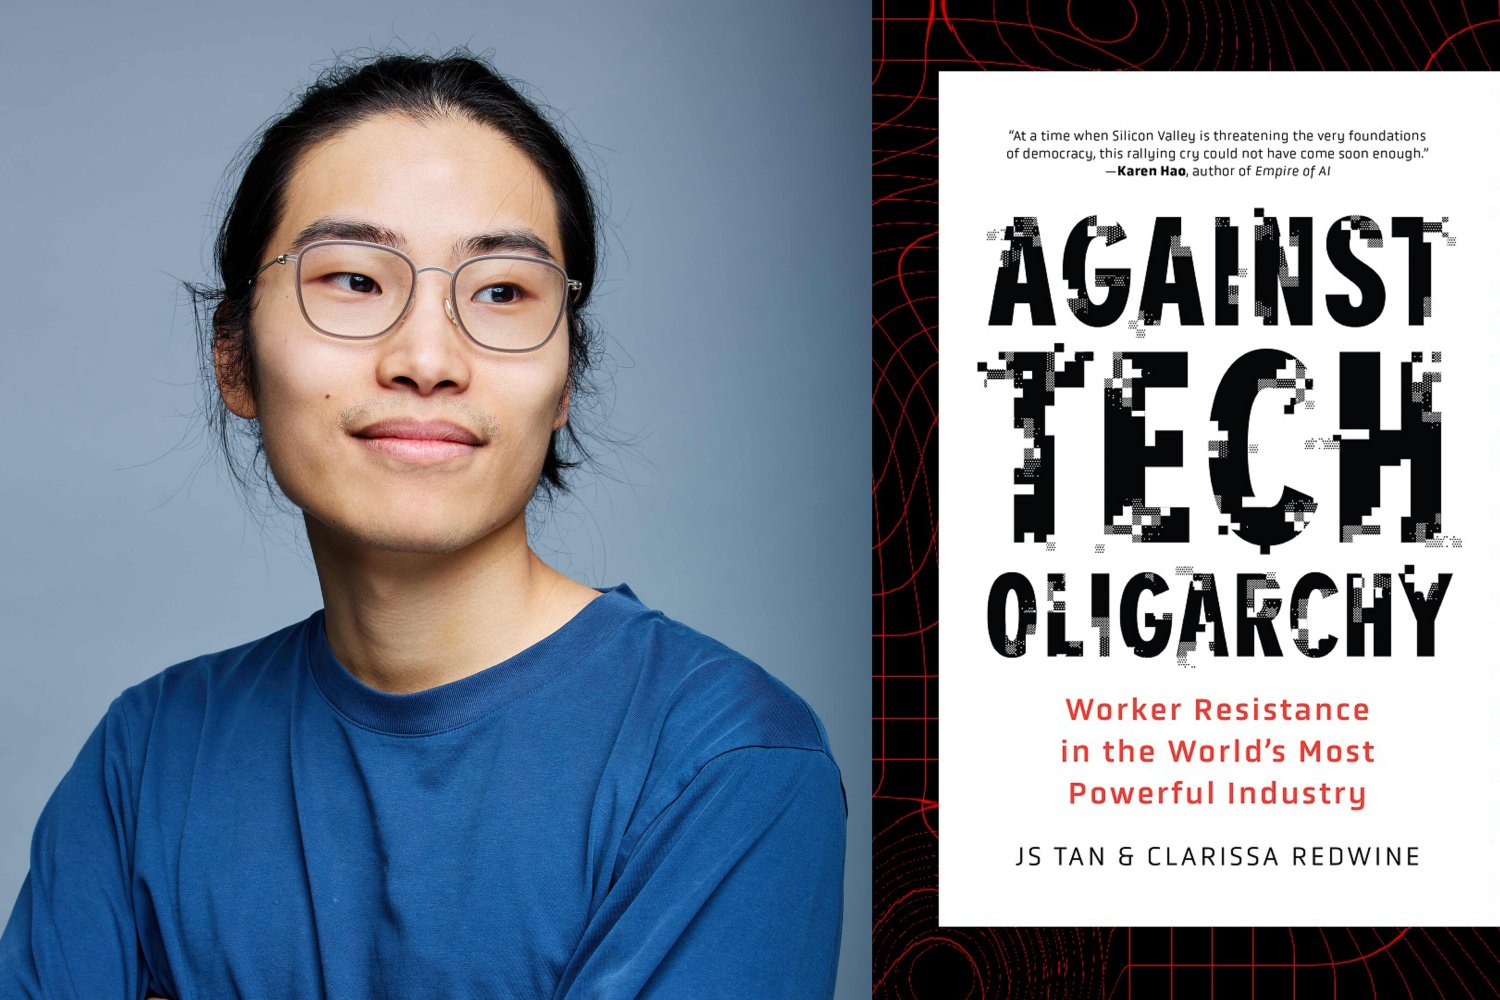
Writing as a participant and researcher, PhD student JS Tan SM ’22 has co-authored a new book about the rise of tech worker protests and the employer backlash that followed.
Requirements in large systems rarely exist in isolation. Their meaning depends on the wider project context - other requirements, policies, decisions, tests, and implementation details. That becomes especially important when AI is used for review, because spotting a possible conflict or gap is only the beginning. ReqSpace explores how AI, visualisation, and connected project context can help reviewers understand those findings, trace the relationships behind them, and focus on the questions that…
AI is making software generation faster, but speed does not remove the need for expertise. As more work is delegated to AI, tacit knowledge may become one of the most important human advantages in software engineering. The post Beyond Prompt Engineering: The Role of Tacit Knowledge in Software Engineering appeared first on GPT-Lab.
We use cookies to run the site and, with your consent, for analytics and to show ads.
See our Cookie Policy.