Skip to content

Data-efficient exploration of atomic clusters via uncertainty quantification on complex potential energy surfaces.

Sep 2026 · Journal of Chemical Physics · Vol 165 9 · 0 citations · 44 references
Medicine

TL;DR

This study developed and evaluated three different UQ frameworks based on advanced UQ methods and integrated with graph neural networks: Bayesian Neural Networks, Evidence Neural Networks, and Monte Carlo Dropout, and shows that ENN effectively reflects data adequacy, while MCD exhibits excellent robustness.

Abstract

Machine learning interatomic potentials have become an effective method for exploring complex potential energy surfaces; however, their application to atomic clusters is frequently hindered by the high cost of sampling diverse isomer spaces and the difficulty in ensuring model generalizability across complex energy landscapes. While uncertainty quantification (UQ) offers a pathway to mitigate data scarcity, its efficacy in capturing continuous potential energy surface features and guiding active learning within the complex landscape of clusters remains systematically unverified. In this study, we developed and evaluated three different UQ frameworks based on advanced UQ methods and integrated with graph neural networks: Bayesian Neural Networks, Evidence Neural Networks (ENN), and Monte Carlo Dropout (MCD). We first validated these models on the MD17 dataset to establish baseline performance, followed by a rigorous assessment on complex cluster systems (Ta2N3- and LaSi24) to probe their decision-making mechanisms in high-dimensional spaces. Our results show that ENN effectively reflects data adequacy, while MCD exhibits excellent robustness, with an MSE of <0.3 eV2 in the first 20 epochs. Furthermore, active learning driven by MCD significantly reduces the computational overhead of first-principles calculations while maintaining high predictive accuracy. This study provides a physics-informed guideline for selecting UQ strategies, facilitating the autonomous and efficient discovery of stable cluster isomers.
View source

Similar papers

Aug 2026

Active Learning for Optimizing Adsorption Energy Predictions in Large Chemical Spaces.

A global active learning framework is demonstrated to map these landscapes efficiently by coupling genetic algorithms with deep neural networks trained on density functional theory data, which provides a scalable and resource-efficient strategy for high-throughput materials discovery in applications such as hydrogen st...

J. von der Heyde, Walter Malone, A. Kara · 0 citations
Open access Sep 2026

Active delta-learning for surface hopping: efficient fitting of potential energy surfaces.

This work combines active learning with Δ-learning - where the ML model predicts the difference between a target quantum chemical method and a computationally efficient baseline - into an end-to-end protocol for active delta-learning (ADL) of potential energy surfaces for trajectory surface hopping dynamics.

M. Martyka, Joanna Jankowska, Pavlo O. Dral · 0 citations
Aug 2026

Δ -Machine Learning for the Prediction of Metal Complex Properties.

An adaptation of the Δ-ML strategy for quantum property prediction of transition metal complexes is presented, which consistently achieves higher accuracy in the prediction of high-fidelity targets, while demonstrating improved data efficiency and out-of-domain transferability.

Hannes Kneiding, David Balcells · 0 citations
Preprint Sep 2026

Benchmarking Machine-Learning Interatomic Potentials for Dynamical Stability in Inorganic Semiconductor Nanocrystals: A CdSe Case Study

Machine-learning interatomic potentials (MLIPs) enable nanosecond-scale atomistic simulations of inorganic semiconductor nanocrystals, but low errors on held-out configurations do not necessarily guarantee stable molecular dynamics. We benchmark five graph-neural-network MLIPs, SchNet, PaiNN, NequIP, Allegro and MACE,...

Muhmmad Usman, M. Suleymanova, Zain Ul Abideen et al. · 0 citations
Open access Sep 2026

Symbolic ensemble learning enables discovery of fast accurate physics-based interatomic potentials

A hybrid symbolic-neural framework is introduced that unifies the interpretability of the Embedded Atom Method with the adaptability of data-driven learning, and derives a composite potential that surpasses the fidelity of its constituent models.

Bilvin Varughese, Aditya Koneru, Adil Muhammad et al. · 0 citations
Open access Aug 2026

Exploring the chemical space of transition-metal cluster thermodynamics via automated first-principles calculations and machine learning

Atomic clusters serve as the embryos of materials, yet their enormous and compositionally complex chemical space has long hindered systematic exploration of thermodynamic stability. Here, we show that a unified first-principles and machine-learning framework enables large-scale mapping of transition metal cluster therm...

Ning-Zheng Li, Zi-Yue Wang, Zi-Yu Li et al. · 0 citations

Related blog posts

Microsoft Research Blog Jul 13, 2026

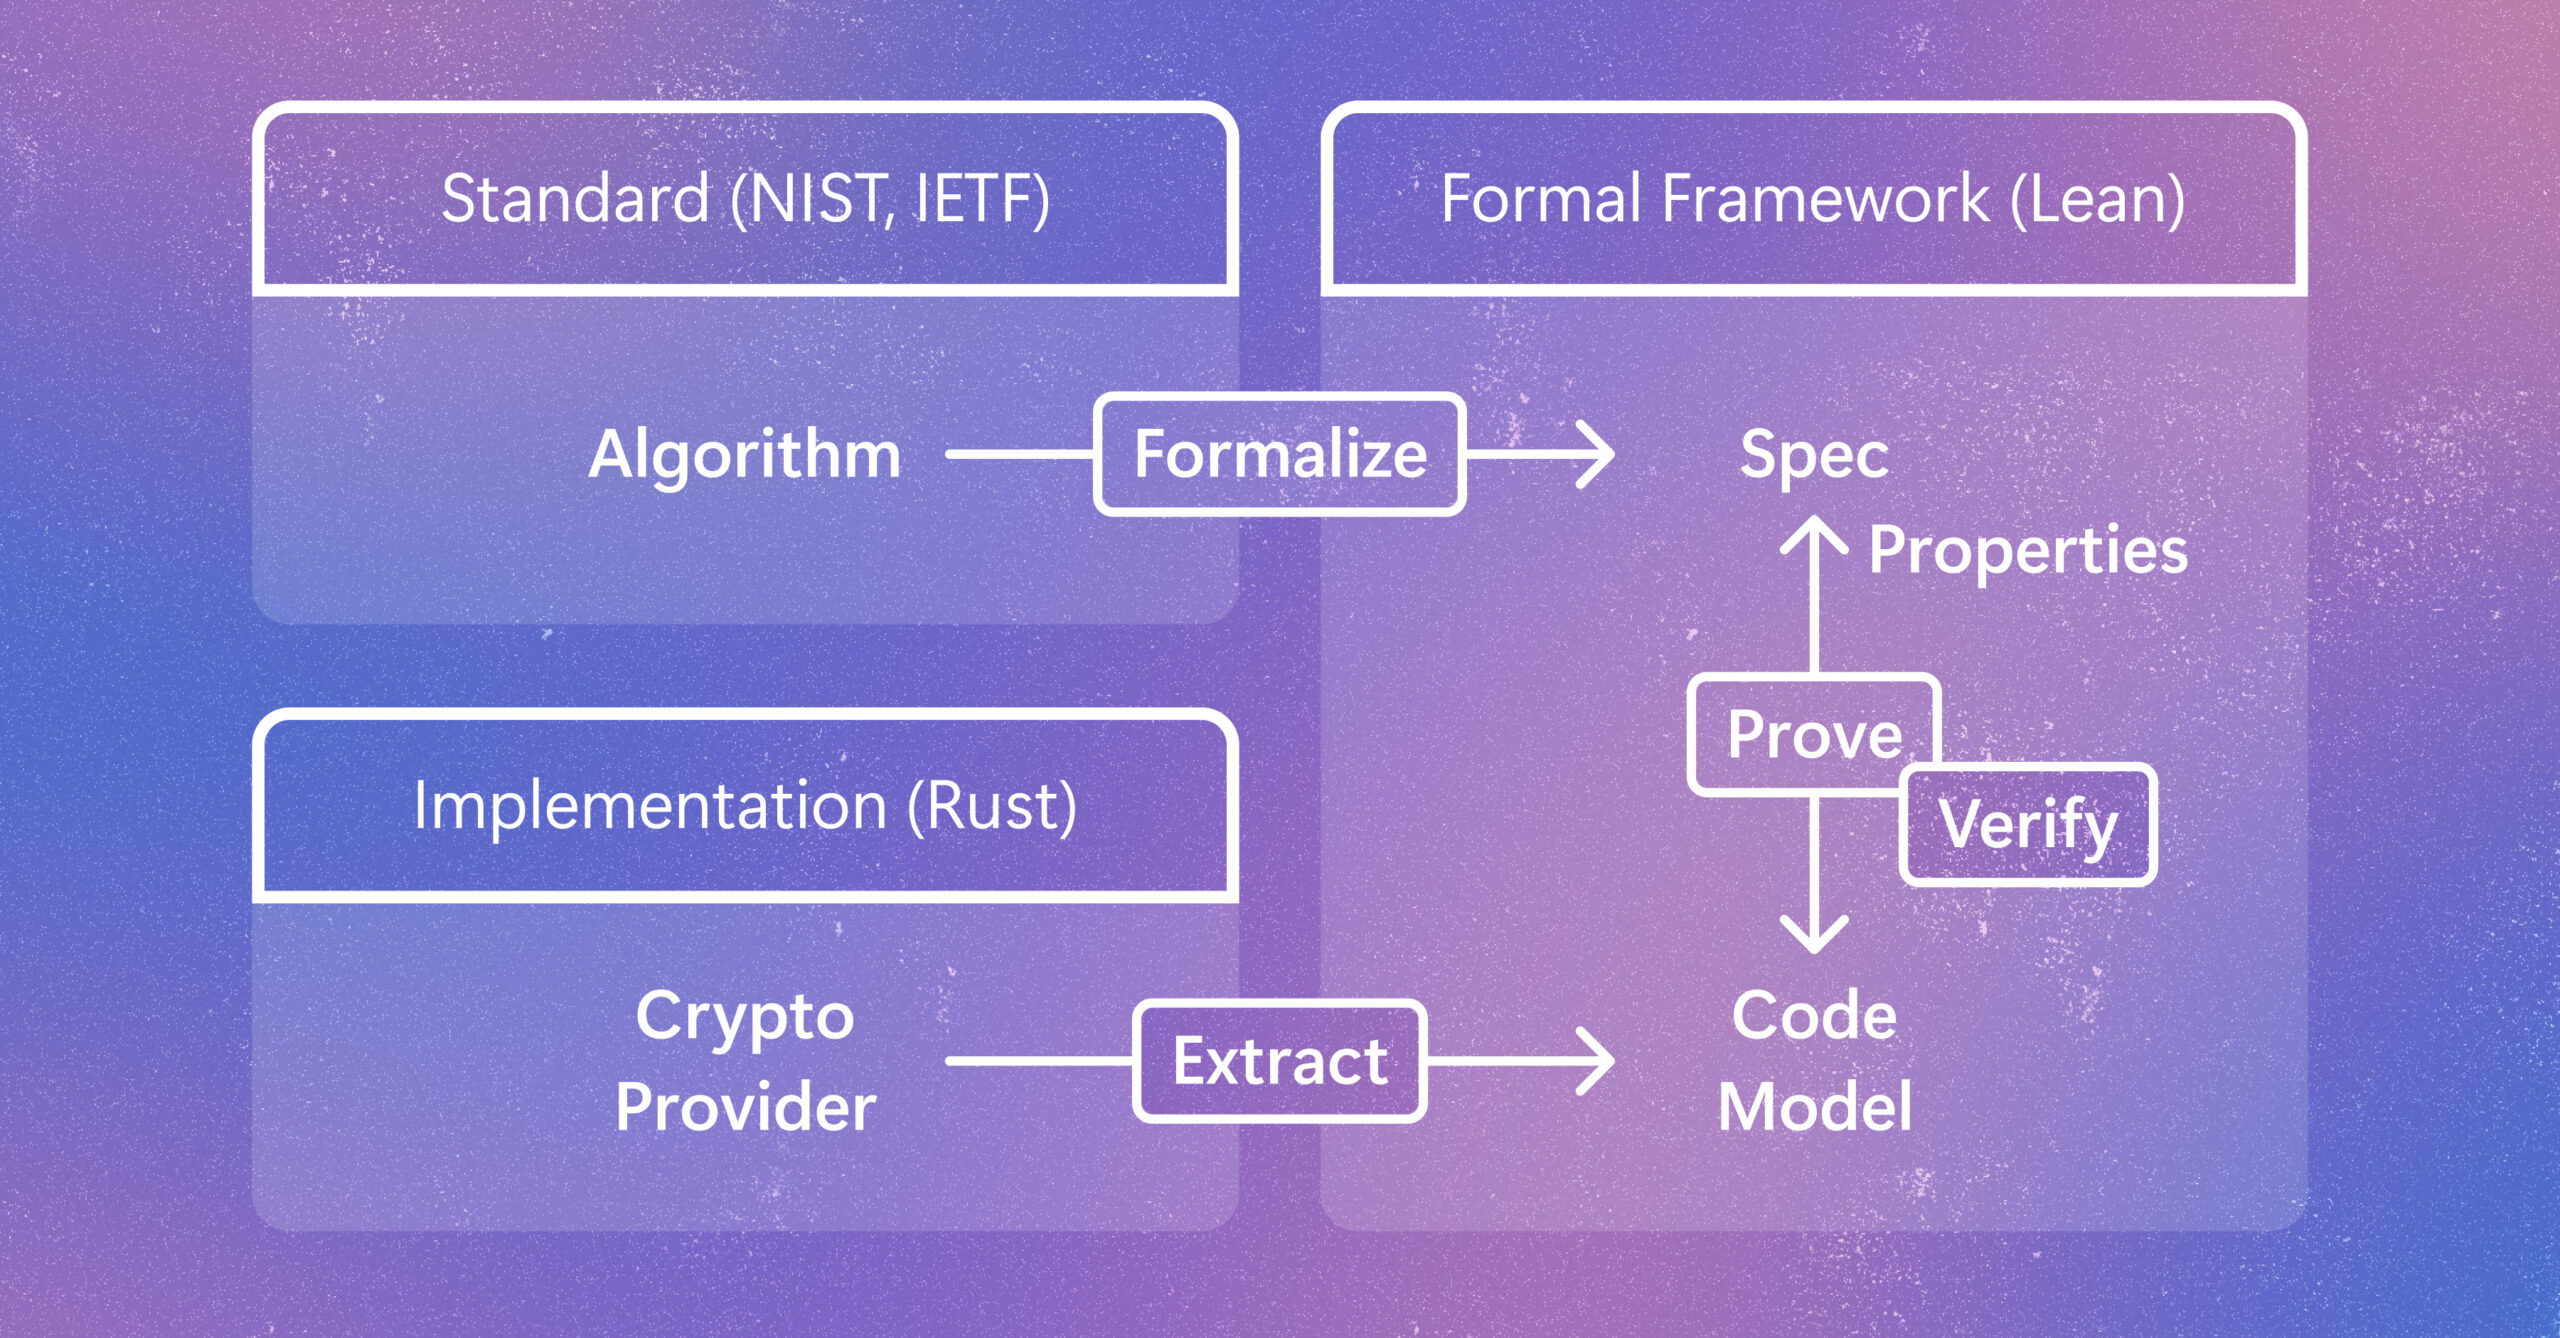
Verifying Rust cryptography in SymCrypt, from standards to code

Cryptographic code supports vital protections in modern computing systems. Learn how a new method helps verify code as developers write it while preserving speed and adaptability as it gets implemented and evolves. The post Verifying Rust cryptography in SymCrypt, from standards to code appeared first on Microsoft Research.

We use cookies to run the site and, with your consent, for analytics and to show ads. See our Cookie Policy.