Rational Coformer Screening of Multicomponent Crystals of Abiraterone Acetate Using Machine Learning (ML) and Multicomponent Hydrogen Bond Propensity (MCHBP): A Comprehensive Structural and Computational Investigation
Sep 2026· Crystal Growth & Design· Vol 26, pp. 7667-7682· 0 citations· 69 references
Machine Learning in Materials ScienceComputational Drug Discovery Methods
TL;DR
A novel predictive framework that combines machine learning with molecular complementarity hydrogen-bond propensity (MCHBP) for efficient screening of coformers for efficient screening of coformers is developed.
Abstract
Multicomponent crystalline systems offer a viable approach to improving the physicochemical and pharmacokinetic properties of drugs, thereby enabling the rational optimization of solid forms through coformer identification. The conventional process relies on empirical knowledge and chemical intuition. To overcome traditional challenges, we developed a novel predictive framework that combines machine learning (ML) with molecular complementarity hydrogen-bond propensity (MCHBP) for efficient screening of coformers. The effectiveness of the method was confirmed in a proof-of-concept study of the pharmaceutical multicomponent system for abiraterone acetate (AA). The success of this process lies in the generation of high-quality three-dimensional molecular descriptors derived from accurate structures. The novelty of this study is that it uniquely obtains high-quality 3D descriptors using optimized structures with the MMFF94 force field. For each molecule, 1,826 three-dimensional Mordred descriptors were generated with RDKit software. We used two types of descriptor fusion strategies: concatenation (M1) and summation (M2). The performance of the ten machine learning models was evaluated; gradient boosting performed well among the other nine tested models. The predicted API-coformer pairs were ranked in descending order of their probabilities. However, the predicted probabilities of the top 20 candidates were very close, making further prioritization difficult. To refine the selection, we used the MCHBP ranking, which enhanced the matching of hydrogen-bond networks. Following the MCHBP analysis, we selected the top eight unreported coformers for experimental investigation. From these, seven new multicomponent phases were identified, and six multicomponent crystals of AA were confirmed through SCXRD: AA−2HBA, AA−23DHBA, and AA−24DHBA as cocrystals and AA−25DHBA, AA−26DHBA, and AA−246THBA·H2O as salts. The type and nature of intermolecular interactions and their contributions were analyzed by Hirshfeld surface and fingerprint analyses. The electronic structure, charge transfer, and hydrogen bonding were analyzed by density functional theory at the DFT/B3LYP/6-311++G(d,p) level. The cocrystal and salt forms were differentiated using the quantum theory of atoms in molecules (QTAIM) and natural bonding orbital (NBO) analysis. Noncovalent interactions were analyzed by noncovalent interaction (NCI) and reduced density gradient (RDG) graphical user interfaces.
Overall, 5f was computationally prioritized based on its favorable predicted EGFR interactions and sustained association over the investigated simulation timescale, while its physicochemical and pharmacokinetic limitations indicate the need for further structural optimization and experimental validation.
Arleta Rifati-Nixha, Mustafa Arslan, M. Dërmaku-Sopjani et al.· Molecules· 0 citations
An alchemical free-energy framework that incorporates peptide-membrane affinity calculations into sequence reoptimization and establishes a physically grounded framework that integrates empirical AMP redesign with free-energy-based thermodynamic evaluation of multiresidue reoptimization.
Chun-Suo Tian, Meng-Chen Zhou, Hai-Peng Wang et al.· Journal of Chemical Theory a...· 0 citations
The electrochemical CO reduction reaction (CORR) by single-atom catalysts (SACs) provides a promising route for converting CO into value-added C1 products, but the rational discovery of efficient SACs remains challenging because of the large configurational space and complex reaction network. Herein, we chose two-dim...
Zi-Yue Zhang, Meng-Meng Xu, Juan Xie et al.· Journal of Physical Chemistr...· 0 citations
An optimized 3D-MoRSE (OPT3D) molecular representation is developed and evaluated that introduces a tunable distance-scale factor to better capture informative interatomic distance regimes and exhibits strong robustness under moderate noise.
Xiao-Peng Li, Zhou-Fa Hu, Ning-Yi Cui et al.· npj Computational Materials· 0 citations
ChalcoVina is presented, an augmented molecular docking program based on AutoDock Vina that simultaneously evaluates intermolecular and intramolecular ChBs for the first time and can be a blueprint for other σ-hole-mediated interactions, facilitating the inclusion of more nonclassical interactions in SBDD.
Wen-Hao Cai, Zi-Yue Li, Yi-Jie Wang et al.· Journal of Chemical Informat...· 0 citations
Related blog posts
Microsoft Research Blog· microsoft.comNov 12, 2026
What's new in research at Microsoft? Host Anson Ho explores the latest, including AI-powered universal translation, a breakthrough in a 31-year-old mathematics challenge, and advances making AI coding agents faster and more efficient. The post Microsoft Research Focus – Edition #98 appeared first on Microsoft Research.
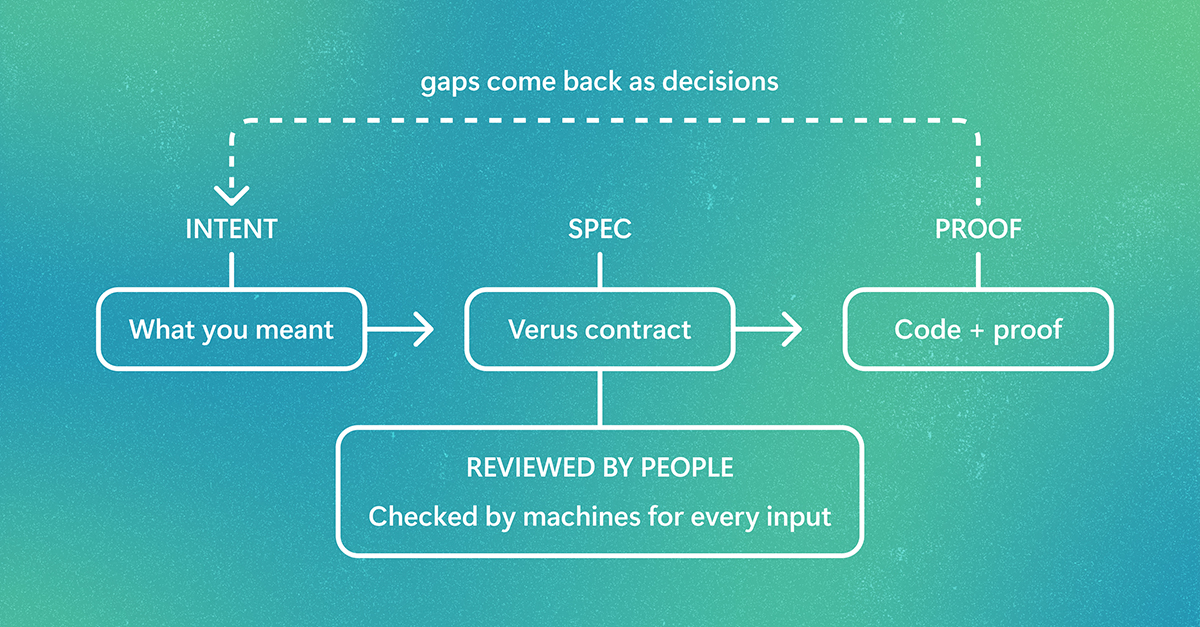
AI agents can generate code at a scale that’s hard for humans to review line-by-line. PDD Kit helps ensure the code is doing what the developer intended by creating formal specifications and testing against them. It converts developer intent into a Verus specification, proves AI-generated code meets it for every input, and uses tests where proofs aren't practical. Developers review intent, not every line. The post Formally verifying agent-written full-stack web apps with PDD Kit appeared first o…