Skip to content
#protein folding Review Open access

From Static Structures to Conformational Ensembles: MSA Perturbation Strategies for AlphaFold2

Oct 2026 · Theoretical and Natural Science · 0 citations
Protein Structure and Dynamics

TL;DR

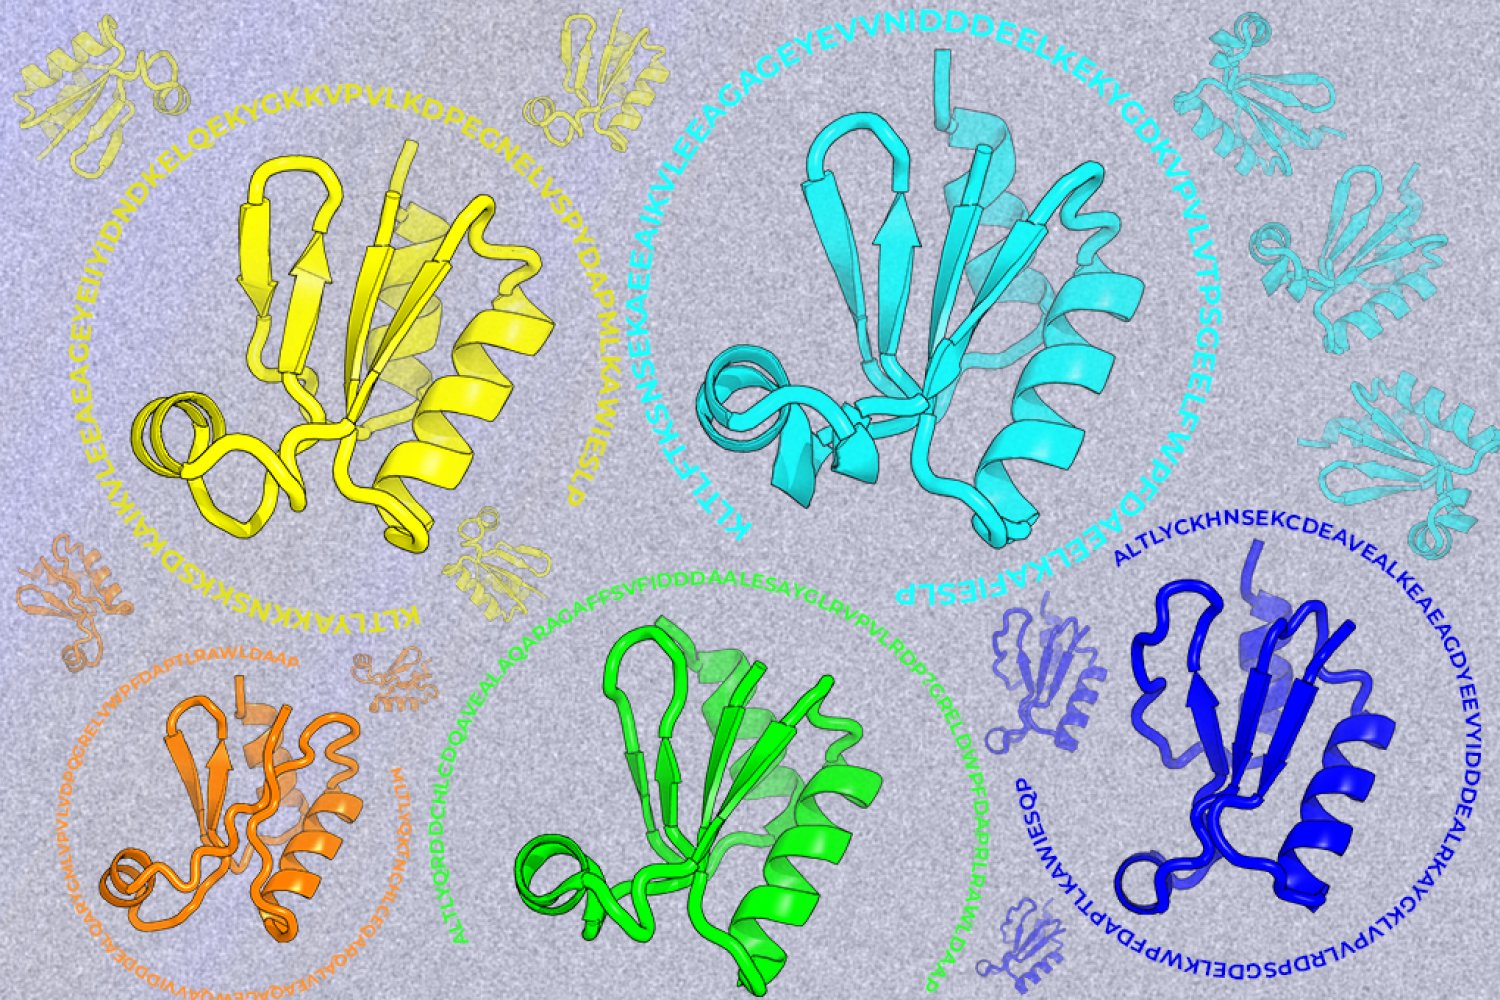
This review synthesizes three complementary strategies for extending AF2 beyond a static prediction by perturbing the multiple sequence alignment (MSA) it receives: sequence-similarity clustering of MSA rows (AF-Cluster), stochastic subsampling of MSA depth combined with dropout and repeated inference, and random masking of MSA columns (AFsample2).

Abstract

AlphaFold2 (AF2) predicts a protein's most probable fold with remarkable accuracy, but native proteins function as ensembles of interconverting conformations, and a single confident output can conceal the very heterogeneity that governs regulation and disease. This review synthesizes three complementary strategies for extending AF2 beyond a static prediction by perturbing the multiple sequence alignment (MSA) it receives: sequence-similarity clustering of MSA rows (AF-Cluster), stochastic subsampling of MSA depth combined with dropout and repeated inference, and random masking of MSA columns (AFsample2). Read together, these methods trace a progression from discovering hidden folds, to comparing relative state populations across mutants and homologs, to generating large, benchmarked ensembles across open-closed proteins and membrane transporters. Reported successes include recovery of both experimentally validated states for a KaiB variant confirmed by NMR, a designed triple mutant whose fold-switched population rose from 11% to 86%, correct directional prediction of population shifts for more than 80% of tested Abl1 kinase variants, and recovery of both reference conformations above a TM-score of 0.8 for 78.3% of OC23 benchmark targets, compared with 47.8% for standard AF2. At the same time, a growing body of independent critique complicates any straightforward reading of these results as evidence that AF2 has learned protein energetics. This review integrates that debate with the original studies' own caveats about pLDDT, MSA composition, and benchmark design to argue that MSA perturbation has made AF2 a valuable generator of testable conformational hypotheses, but not yet a validated or calibrated model of the protein conformational ensemble.

Read PDF

Similar papers

Open access Apr 2026

Several multiple sequence alignment-perturbing methods enhance AlphaFold3 sampling of alternative protein states

Protein function often involves multiple conformational states. Several multiple sequence alignment-perturbing strategies, including stochastic subsampling, clustering, and column masking, have been shown to enhance AlphaFold2 (AF2) sampling of alternative protein states. Here, we evaluate these strategies on AlphaFold...

Samuel Eriksson Lidbrink, Ivan Nissen, Jonathan Kenichi Ahrlind et al. · 0 citations
Open access Oct 2026

Predicting Alternative Protein Conformations with AlphaFold2

Proteins are dynamic molecules capable of transitioning between open, closed, active, inactive, bound, and unbound conformations in accordance with the environment inside the cell. As a result, a given protein can have one conformation that is easily observable and another one which plays its role in signaling, transpo...

Max Wang · 0 citations

A Shortcut to Structure in AlphaFold 3

AlphaFold 3 predicts protein structures with remarkable accuracy, yet how structural information emerges within the model remains poorly understood. Here, through causal interventions on internal representations and direct probing of every Pairformer block, we trace the formation of global protein geometry and identify...

J. Feldman, J. Skolnick · 1 citation
#protein folding Open access Aug 2026

Accurate and efficient prediction of protein conformations with ProtMonomer

Deep learning-based protein structure prediction methods that leverage evolutionary information from multiple sequence alignments (MSAs), exemplified by AlphaFold2, have achieved remarkable accuracy. However, existing methods still struggle to predict challenging proteins, particularly those with novel folds or limited...

Yunda Si, Suqi Zhang, Luo-Nan Chen · 0 citations
Open access Oct 2026

Mapping Functional Transitions across Protein Homologues from a Single Simulation: Application of gEDES-HM to Adenylate Kinases

GEDES-HM (generalized Ensemble Docking with Enhanced sampling of pocket Shape-Homology Modeling), a scalable computational protocol enabling us to remap conformational ensembles obtained from enhanced sampling simulations of a single protein to its homologous family members, removing the need to perform individual simu...

Han Kurt, Mohd Athar, Andrea Basciu et al. · 0 citations

Related blog posts

Google DeepMind Blog Sep 30, 2026

Introducing SynthID Bio

Proof of concept for watermarking AI-generated proteins while preserving biological function.

MIT News · Artificial Intelligence Aug 27, 2026

Looking beyond natural sequences

A new machine-learning framework aims to improve the success rate of computational protein design while moving away from results that reproduce sequences found in nature.

We use cookies to run the site and, with your consent, for analytics and to show ads. See our Cookie Policy.