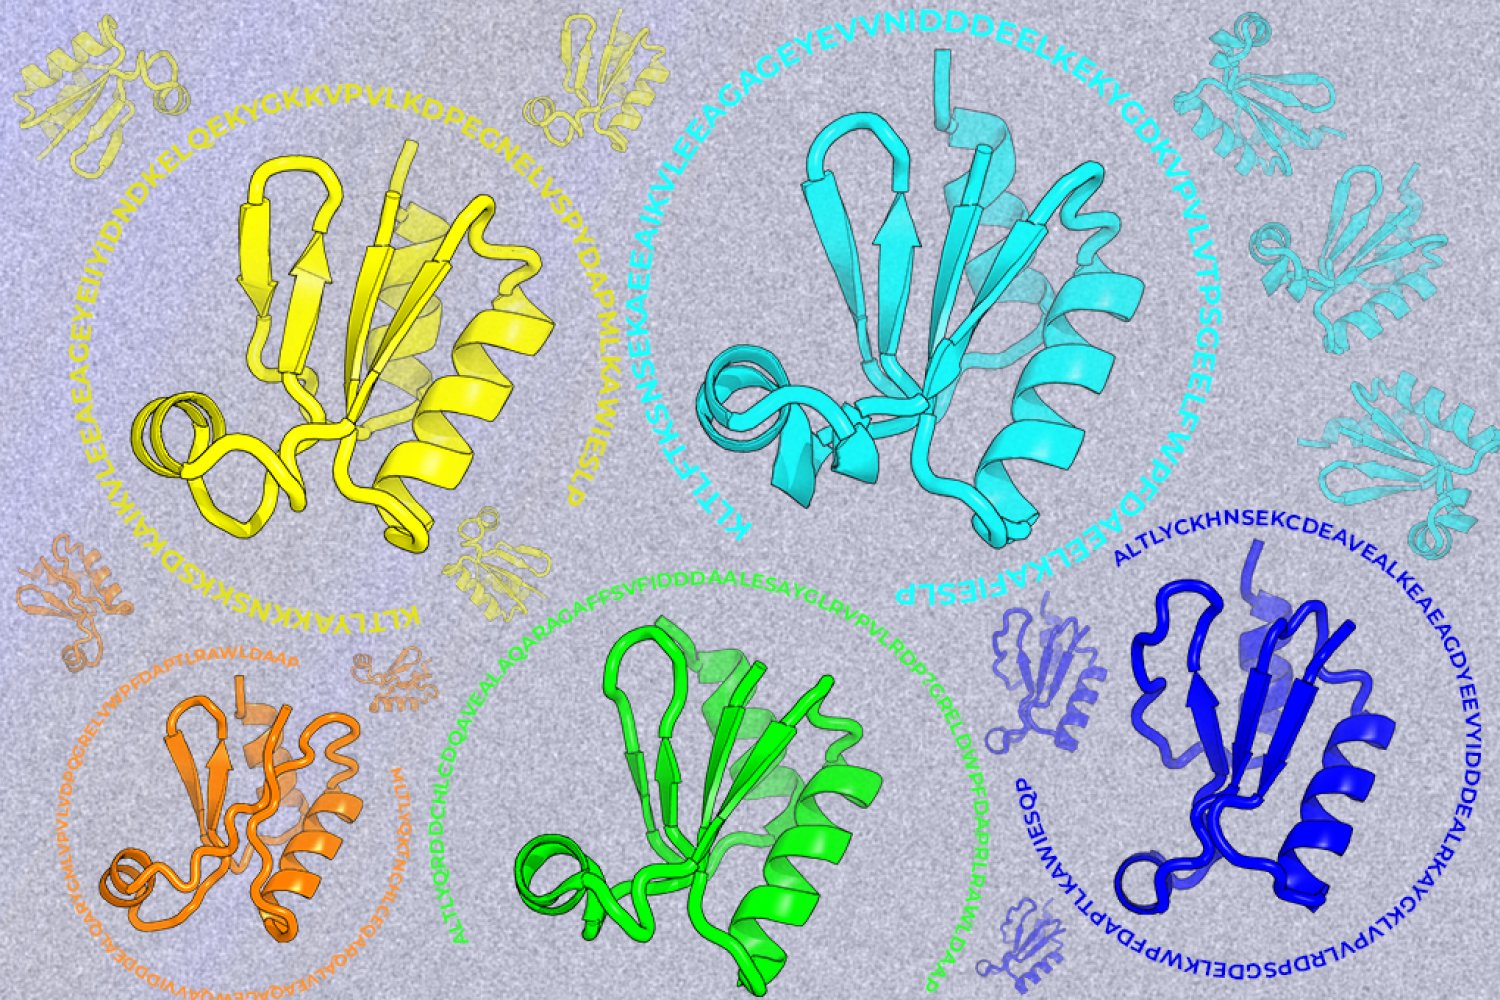
High resolution LATS2:YAPWW1 crystal structure and molecular dynamics simulations suggest that preorganized peptides bind more tightly and less selectively, while flexible peptides incur larger binding penalties that dynamic YAPWW1 compensate more effectively.
Abstract
The two paralogs YAP and TAZ act through TEAD transcription factors and bind PPxY motif proteins in the Hippo pathway via WW domains. YAP has up to two WW domains, whereas TAZ has one. Because YAPWW1 and TAZWW are the most similar in sequence, they can be considered as corresponding modules in these two proteins. This study shows that, despite their similarity, they differ strongly in conformational stability and in how they bind flexible ligands. Nano-differential scanning fluorimetry, circular dichroism, and NMR indicate that both isolated domains populate partially folded or exchanging states in solution, but TAZWW is more thermally stable and has a larger folded population. Peptide binding stabilizes both domains, producing sharper NMR signals. Surface plasmon resonance measurements with PPxY peptides show micromolar affinities and generally modest differences between YAPWW1 and TAZWW, although these differences increase for conformationally plastic ligands. A high resolution LATS2:YAPWW1 crystal structure and molecular dynamics simulations suggest that preorganized peptides bind more tightly and less selectively, while flexible peptides incur larger binding penalties that dynamic YAPWW1 compensates more effectively. These observations indicate that differences in WW-domain stability contribute to subtle YAP/TAZ binding preferences despite conserved binding surfaces and evolutionary relatedness between the paralogs.
Background: Ephrin type-A receptor 1 (EphA1) receptor tyrosine kinase plays a pivotal role in disease development by orchestrating cell-cell communication. Although phosphorylation regulates Eph receptor signaling, its dynamic impact on sterile alpha motif (SAM) domain conformation and function remains poorly understoo...
Madhura Mundhe, Vishakha Kamble, Ishwari H. Sidwadkar· Journal of Integrative and T...· 0 citations
The data support previously reported sequence hotspots, whereby certain PBMs are recognized by large numbers of PDZ domains, likely due to a combination of widely favorable modulator residues, and support previously reported sequence hotspots.
Cole D. Masuga, Caroline Ceravolo, Elise F. Tahti et al.· Protein Science· 0 citations
Aminopeptidase N (APN) is a critical exopeptidase that plays key roles in physiological processes such as tumor metastasis, immune regulation, and viral infections. However, the conformational dynamics associated with different APN inhibitors remain insufficiently understood, particularly whether inhibitors with varyin...
Shuang Yu, Jin-Hong Wang, Yi Zhao et al.· Journal of Molecular Graphic...· 0 citations
Protein phosphatases counterbalance kinases by dephosphorylating phospho-proteins to regulate signaling pathways. But unlike kinases, their shallow catalytic groove has limited selectivity for the phospho-peptide motifs. Phosphatases such as protein phosphatase-1 (PP1) recognize their substrates through short linear mo...
Evi Setiani Lande, M. Kjaergaard· bioRxiv· 0 citations
SAMD9 and SAMD9L (SAMD9/9L) are large cytosolic proteins essential for hematopoietic homeostasis and antiviral defense (1–3). Germline gain-of-function (GoF) mutations in SAMD9/9L cause severe multisystem disorders and predispose to leukemia, but the mechanisms that regulate SAMD9/9L activity and how pathogenic mutatio...
Z. Mou, Fu-Shun Zhang, Marisol Morales et al.· Science Advances· 0 citations
SUMMARY Pseudokinase domains (PsKDs) mediate key signaling processes involved in human disease. Despite transformative success in targeting catalytically active protein kinases, drug development for pseudokinases has lagged behind. Although they lack catalytic activity, pseudokinases are conformationally dynamic and ar...
Joshua B. Sheetz, Yuko Tsutsui, Kumar D. Ashtekar et al.· Structure· 0 citations
A new machine-learning framework aims to improve the success rate of computational protein design while moving away from results that reproduce sequences found in nature.